When a Laboratory Certificate Is Not Enough: Why Research Material Traceability, Regulatory Status and Supplier Documentation Are Three Different Questions
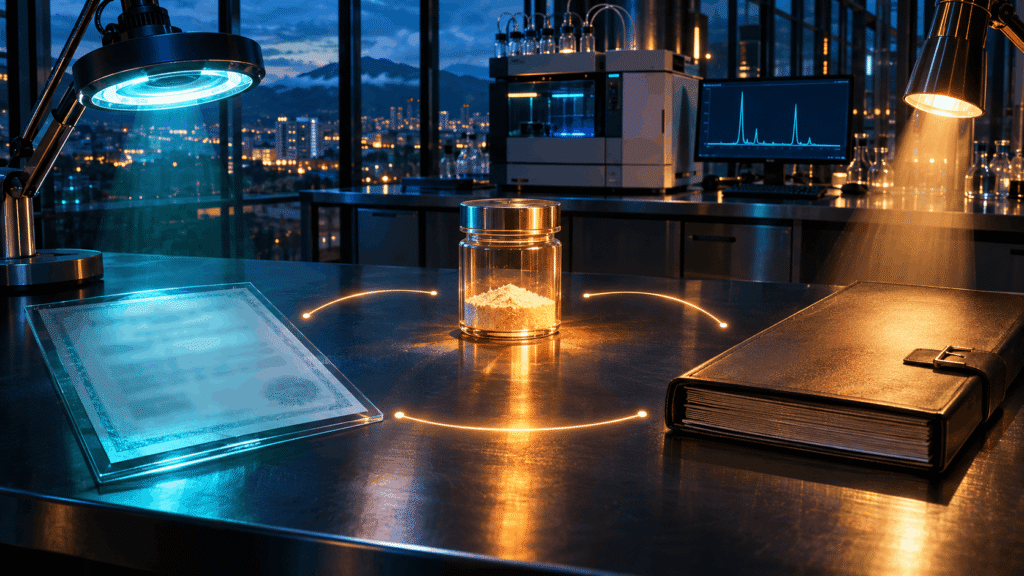
A laboratory receives a certificate of analysis showing a reported purity result for a research material. The document identifies a testing method and includes a date, a laboratory name, and a reference number. To someone looking only at the headline result, the paperwork may appear to answer the most important question: whether the material has been tested.
But the certificate does not necessarily establish which physical shipment the result applies to, whether the receiving organization possesses the same lot that was analyzed, or whether the material can legally be marketed for a particular purpose in the country where it is offered.
These are three different questions. Analytical testing concerns the characteristics measured in a sample. Traceability concerns the connection between the sample, the test result, and the material later handled by a laboratory. Regulatory authorization concerns the legal status of a particular product and its intended use within an applicable jurisdiction.
Research organizations that treat these questions as interchangeable can end up with documentation that looks complete while important uncertainties remain unresolved. A reliable procurement process needs to distinguish each type of evidence and recognize what it cannot prove.
A certificate of analysis is evidence of a test, not proof of everything about a product
A certificate of analysis, often abbreviated COA, is a document that reports analytical findings for a sample or batch. Depending on the material and testing purpose, it may contain information about identity, purity, impurities, composition, or other properties.
The usefulness of the certificate depends on what was tested, which method was used, how the results were interpreted, and whether the report can be connected to the material under consideration.
For example, a chromatographic purity percentage may describe how detected components were represented under a specified analytical method. It does not necessarily establish the total amount of the intended compound in the sample, the absence of every possible contaminant, or suitability for a use that was never evaluated.
Similarly, a mass spectrometric result may support a conclusion about molecular identity without independently proving all aspects of sample quality.
Those distinctions are not weaknesses unique to one analytical technique. All scientific measurements have defined purposes and limitations. Good reporting explains the scope of the result instead of expecting one number to answer unrelated questions.
The first practical step is therefore to ask what the certificate actually supports. If the test concerns chemical identity, it should be read as evidence relevant to identity. If the report includes a purity measurement, the result should be interpreted in the context of its method and the particular material tested.
Traceability answers the question that a purity percentage cannot
Imagine that a research organization receives two separate shipments of the same nominal compound. Both packages carry the same general product description, but each has a different lot identifier.
The first lot has an analytical report, while the second arrives with a file whose name contains only the product description and testing date. A technician stores both documents in a shared folder. Months later, another researcher attempts to establish which report supports the material used in a published experiment.
If the organization did not preserve the relationship between the shipment, lot, internal inventory identifier, and analytical report, the presence of the certificate may not resolve the uncertainty.
This is a documentation and traceability problem rather than a dispute over the analytical method.
Research information pages such as the Tirzepatida Costa Rica research information overview describe the role of technical documentation and lot records in the research-material supply process. Such descriptions can help readers identify what documentation to request, but the underlying records still require independent review.
A traceable laboratory record should make it possible to establish which material was received, how it was identified, what supporting documentation accompanied it, and how that material was connected to subsequent analytical or experimental work.
Where identifiers do not match, the discrepancy should be investigated instead of resolved through an undocumented assumption.
Why laboratories need more than a supplier’s product name
Product names are designed to help people recognize materials, but they may be insufficient for rigorous scientific recordkeeping.
One supplier may use a descriptive commercial label, while another uses a chemical name, research code, or abbreviated sequence. Different testing laboratories may assign their own sample identifiers to the same submitted material.
The resulting documents can refer to one substance using several different names, even when every participant has recorded information accurately within its own system.
To maintain traceability, a research organization may need to preserve multiple identifiers and record how they relate to each other.
That includes distinguishing a general compound designation from the identity of a particular batch. Two batches associated with the same chemical name should not automatically be assumed to possess identical measured characteristics.
The relevant documentation should also indicate whether analytical testing was performed on the batch in question or on another sample intended to represent it.
Without those distinctions, an apparently complete report can be interpreted more broadly than the evidence allows.
Regulatory authorization is a separate question from analytical quality
A laboratory report can establish information about a sample without determining whether the associated product is authorized for marketing or a particular use.
Regulatory status depends on the applicable national rules, the nature of the product, the claims made about it, its intended use, and any relevant registration or authorization requirements.
The distinction is especially important when scientific terminology appears alongside commercial offers for compounds also associated with pharmaceutical research.
In Costa Rica, the Ministerio de Salud issued a sanitary warning in July 2026 and a follow-up communication on September 14, 2026, concerning the promotion and commercialization of certain products containing or claiming to contain substances such as semaglutide and tirzepatide.
The September communication identified concerns involving products without current sanitary registration and insufficient information to establish their origin, composition, manufacturing conditions, storage, or distribution.
These official warnings provide regulatory context. They do not automatically determine the legal status of every research supplier or every product described on the internet.
For a specific product, legal status should be established using current official records and the applicable legal classification. An analytical certificate should not be treated as a replacement for that review.
The research-use label has a limited meaning
Research-oriented material descriptions often state that a compound is intended exclusively for laboratory investigation.
Such language communicates a stated intended use. It does not, by itself, prove that every aspect of the product’s manufacture, distribution, marketing, or importation satisfies the laws of a particular country.
Nor does a research-use statement establish that the material has been assessed as safe or effective for human treatment.
These distinctions matter because scientific investigation, product availability, and authorization for a medical use are separate stages or categories of activity.
The United States Food and Drug Administration describes drug development as a progression that includes discovery, preclinical research, human clinical research, regulatory review, and subsequent monitoring.
That framework illustrates why a laboratory observation cannot substitute for the human evidence and regulatory review needed to support a specific therapeutic claim. US regulatory requirements are not identical to those in Costa Rica, but the conceptual distinction between experimental evidence and authorization remains important.
Educational content should explain these boundaries without suggesting that a commercial designation automatically settles the legal question.
What ISO/IEC 17025 does and does not establish
Another frequently encountered element in laboratory documentation is accreditation or a reference to ISO/IEC 17025.
The International Organization for Standardization describes ISO/IEC 17025 as a standard relevant to the competence, impartiality, and consistent operation of testing and calibration laboratories.
Accreditation can provide confidence concerning activities within a laboratory’s recognized scope. However, the actual scope matters. A laboratory may be accredited for certain methods or testing activities without that accreditation extending to every possible measurement it performs.
Readers should therefore examine the specific laboratory identity, the stated accreditation, and whether the relevant analytical activity falls within its recognized scope.
It would be inaccurate to interpret a general accreditation claim as universal confirmation of the quality, regulatory status, or suitability of every product associated with a supplier.
Likewise, laboratory accreditation should not be confused with authorization to market a drug or with evidence that a compound produces a particular clinical outcome.
These are different questions requiring different records.
The chain of documentation becomes more fragile when materials change hands
Research materials may pass through several organizations or departments before a test is performed.
A manufacturer may create the original lot record. A testing laboratory may receive a sample under a separate identification code. A distributor may maintain an internal inventory reference. The receiving research institution may assign another identifier when the shipment enters its own system.
Each step creates an opportunity for useful information to become separated from the physical material.
A robust documentation process should preserve the connection between the relevant identifiers and record when responsibility for the material changes.
It should also distinguish the date of analysis from the date of distribution and the date of receipt. These events may occur at different times and should not be treated as equivalent.
When information is missing, a clear exception record can be more useful than an informal attempt to reconstruct the history from emails and file names.
The aim is not to create paperwork for its own sake. It is to allow another qualified person to understand what happened without relying on the memory of whoever originally handled the shipment.
A searchable document database is helpful only when the relationships are reliable
Digital document systems make it easier to store and retrieve analytical records, but a file being searchable does not guarantee its connection to the correct material.
A useful system should allow users to search by meaningful identifiers such as a recognized lot reference, internal sample ID, or report number.
It should also distinguish reports associated with different batches or revised analytical findings.
A folder containing hundreds of certificates may appear organized while remaining difficult to audit if the naming rules are inconsistent or the links to inventory records are incomplete.
Structured metadata can improve this situation. Separately storing the laboratory identity, sample reference, report date, method, and relevant document version allows software to compare records more consistently.
Still, automated matching should not replace human investigation when records conflict. A matching file name is not proof that two samples are the same.
Technology can support traceability, but it cannot create missing evidence retroactively.
How to interpret a supplier’s technical statements without assuming endorsement
Websites discussing laboratory materials may describe quality controls, analytical documentation, packaging practices, or research areas.
These descriptions are useful starting points for questions, not independent verification of every stated practice.
For example, the Tirzepatida Costa Rica research website includes descriptions of research materials, chemical information, and documentation-related claims. A reader should distinguish those statements from independently reviewed analytical records or official regulatory findings.
Where a website claims to provide batch-specific certificates, the relevant question is whether the document for the particular material can actually be examined and matched to the identified batch.
Where a website discusses quality control, the reader should ask which tests were performed, by whom, and under which stated methods.
Where the site refers to research-only materials, readers should not assume this is equivalent to authorization for human use or clearance under national commercial rules.
A transparent explanation of evidence levels helps readers assess information without treating an educational mention as an endorsement of the supplier or its products.
Why laboratory findings and human evidence require different interpretations
Scientific research often begins with mechanistic questions involving cells, molecules, or experimental models.
These investigations can reveal how a compound interacts with a biological target under controlled conditions. They may suggest hypotheses worth further exploration.
However, a biological response in a laboratory model does not necessarily establish a clinically meaningful effect in people.
Human studies have their own requirements concerning participants, study design, outcome measurement, safety monitoring, and interpretation.
The FDA’s public development resources distinguish preclinical studies from clinical trials and emphasize that evidence from early investigation must be evaluated within its appropriate stage.
When online content discusses a compound’s interaction with a receptor or other biological target, readers should identify whether the statement describes a biochemical mechanism, an experimental result, or an established clinical outcome.
Those are not interchangeable claims.
Responsible scientific communication identifies the type of evidence supporting a statement, rather than implying a practical benefit merely because a mechanism is scientifically interesting.
Five questions that separate evidence from assumptions
Research organizations and technical readers can evaluate documentation using a consistent set of questions:
- What exactly was tested? Identify the specific material, sample, or batch rather than relying on a broad product description.
- What does the method measure? Determine whether the reported result concerns identity, purity, composition, or another defined analytical property.
- Can the result be traced? Confirm that the test report can be associated with the relevant physical material and its recorded history.
- What is the applicable legal status? Review official requirements and records for the particular product and intended use in the relevant jurisdiction.
- Does the claimed outcome match the evidence? Separate laboratory observations from human clinical findings and avoid conclusions that extend beyond the study design.
These questions help prevent one form of evidence from being used to answer a different question.
They also create a clearer record when a concern is identified. Instead of broadly labeling a product or document as acceptable or unacceptable, an organization can identify the particular information that is missing or unresolved.
Better research procurement is built on separate, connected checks
The most reliable documentation process does not ask a single certificate to establish analytical quality, traceability, regulatory status, and clinical relevance all at once.
It treats those subjects as separate assessments and connects their supporting records where appropriate.
An analytical report can describe measurements made on a defined sample. Traceability records can help establish which material the report concerns. Official regulatory sources can clarify applicable authorization requirements. Published studies can provide evidence relevant to specific scientific questions.
None of these records automatically substitutes for the others.
For research organizations working across suppliers, testing facilities, and national borders, this distinction is particularly important. Different organizations may maintain different identifiers, use different reporting systems, and operate under different legal frameworks.
Good documentation makes those differences explicit rather than hiding them behind a familiar compound name or an attractive summary result.
The central lesson is simple: a certificate can be technically meaningful and still be insufficient for a broader decision. The strongest process is one that identifies precisely what the certificate establishes, preserves the connection to the correct material, and resolves regulatory questions through the authorities responsible for them.

About Author